From a TP53 variant to an evidence-anchored hypothesis in one session.
Enter any variant and read what is actually measured: the Measured binding card searches the public ChEMBL bioactivity archive live for your specific variant, plus same-residue context, and surfaces published Kd, Ki, IC50 and EC50 values with their assay method and original citation. Cohort frequency, functional screens and cited literature sit alongside. The computational shortlist is a transparent physics screen for cavity fit and ligand efficiency, anchored in measured data and published evidence — docking scores are never presented as binding affinities. Every number auditable, every export reproducible.
More than half of human tumours carry a TP53 mutation. Understanding what each variant breaks, and what could reactivate it, is one of oncology's highest-stakes questions.
Oncyra folds the steps a cancer biologist would otherwise chase across many tools, variant entry, receptor selection, mutation-aware modelling, WT overlay, cavity map, live measured-binding lookup against the public ChEMBL archive for your specific variant plus same-residue context, ligand screening against 3,311 approved drugs and a curated rescuer library, per-residue interaction analysis, evidence-linked rescue shortlisting, orthogonal in-cell validation including a live cell physiology simulation, and a reproducible screen export, into a single continuous session across the 4,137 TP53 DBD missense variants in the catalogue.
The problem, the gap, and what Oncyra actually does about it.
TP53 is mutated in over half of human tumours, yet direct reactivators remain rare and mutation-specific. Oncyra collapses the mutation → structure → pocket → ranked ligand loop into a single transparent session across 4,137 catalogued TP53 DBD missense variants and 3,311 approved drugs, and shows its work at every step.
One mutation, one target, years of setup.
Each p53 hotspot (R175H, R248Q, R273H, Y220C, R282W and beyond) has a distinct destabilisation and pocket signature. Standard pipelines rebuild the structural and screening context from scratch per variant.
Pathogenicity scores stop short of a rescue hypothesis.
General pathogenicity scores classify a variant as damaging but do not identify the disrupted structural motif or the ligand class most likely to reactivate it. Oncyra targets that mechanistic gap.
Mutation-aware, mechanistic, and honest.
Mutation-aware candidate model with WT overlay and cavity map, deterministic physics-first screening against 3,311 approved drugs and a curated rescuer library (cavity-fit prefilter, then docking, then ligand-efficiency ranking), curated motif attribution, calibrated uncertainty bands, and DMS overlays, all exported as JSON across the 4,137 catalogued TP53 DBD missense variants.
From mutation to rescue candidate
Five computational phases, presented in the workbench as eleven interface steps. Pick a phase to see its steps and what it hands to the next one.
Ingestion and eligibility
Key output
Eligibility gate: truncating, splice and outside-DBD (94–312) variants stop here
01Variant entry
Enter a TP53 protein change or upload a catalogue drawn from the 4,137 TP53 DBD missense variants on file, of which 884 pass the structural dockability gate. Truncations, splice variants and residues outside the folded DNA-binding domain (94-312) are refused before any fold or dock starts, so the rest of the session works on a well-defined set.
02Receptor selection
Choose an experimental PDB, upload a private PDB or CIF, or use a mutation-aware candidate model when no experimental structure fits the variant.
The platform in action
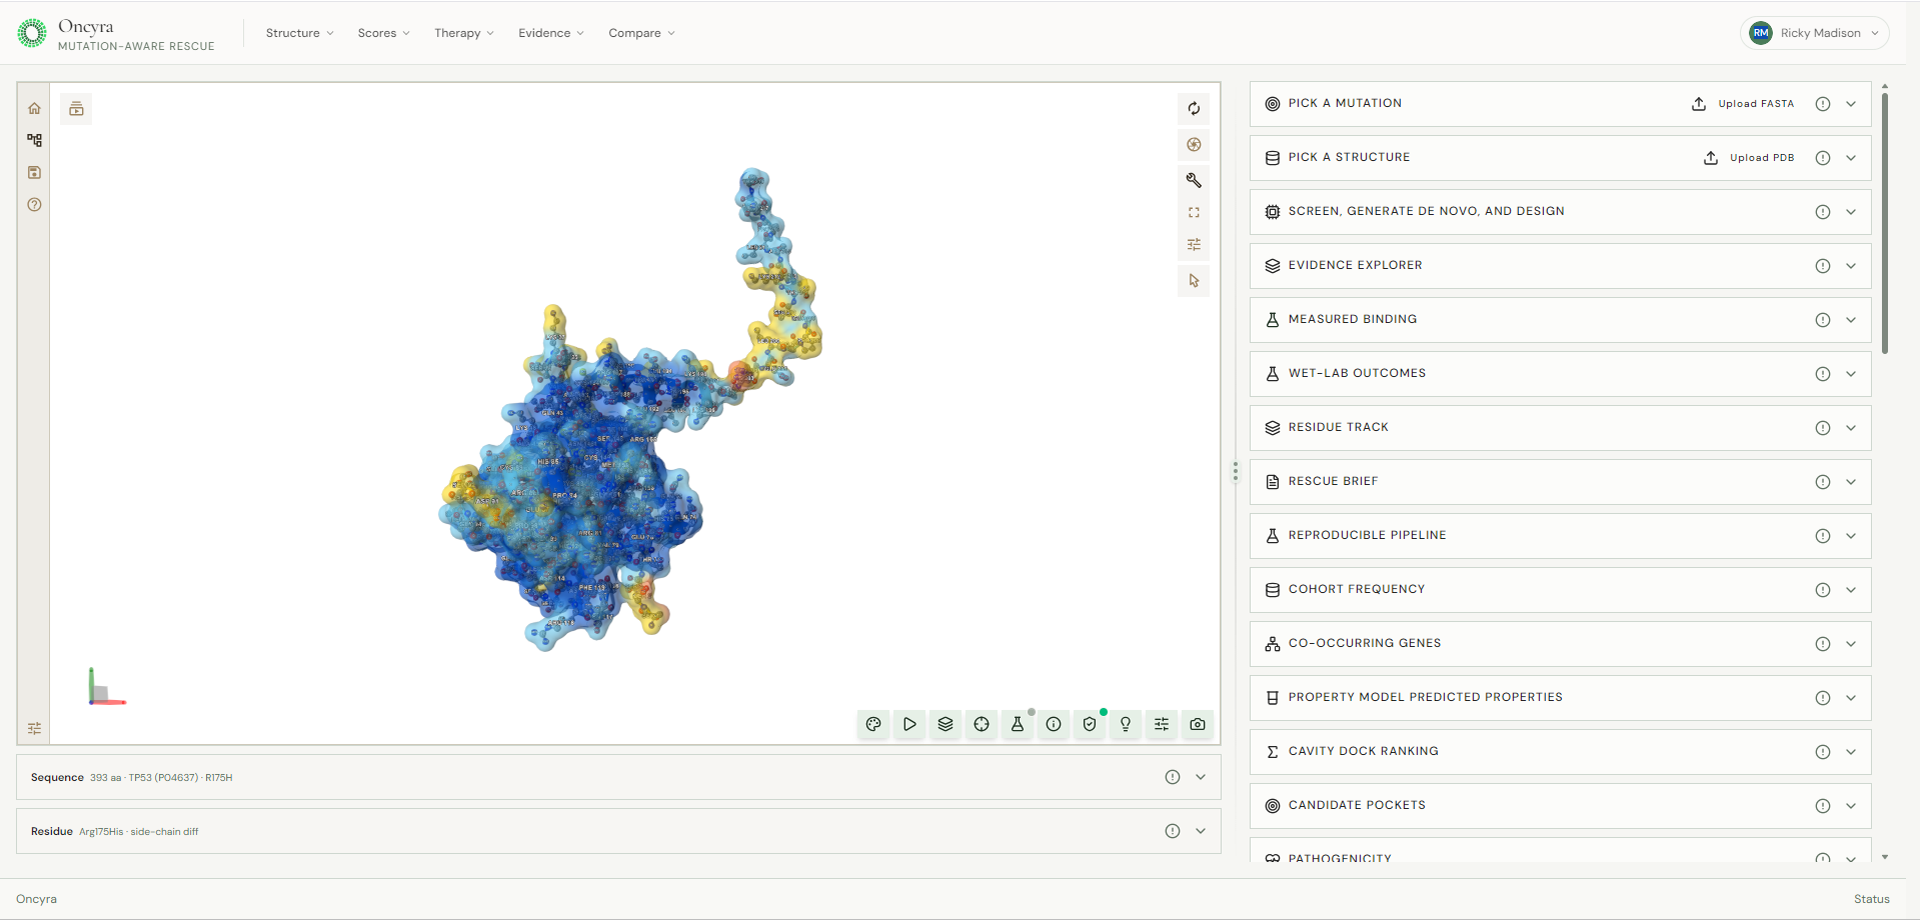
One workbench instead of a chain of tools
Structure predictors and docking engines are excellent at their one job. Oncyra connects them to TP53 genetics, evidence and checks, so a result arrives with its context.
| Capability | Oncyra | AlphaFold / ESMFold | AutoDock Vina | Biohub-style bio-AI models |
|---|---|---|---|---|
| Workflow and rigour | ||||
3D structure of the mutant ESMFold default, opt-in MSA + OpenFold3 | ● | ● | ○ | ◐ |
Blocks ineligible variants before compute Truncating, splice and outside-DBD (94–312) variants are stopped | ● | ○ | ○ | ○ |
Pocket detection and receptor QC In-browser P2Rank port, completeness gate | ● | ○ | ○ | ◐ |
Docking in the same session Local physics, consensus and ensemble passes | ● | ○ | ● | ○ |
Eight pose checks on every result Clashing poses withhold their energy | ● | ○ | ◐ | ○ |
| Interactive chemistry | ||||
Sketch a molecule and dock it in the browser Hand-drawn design lane beside screening | ● | ○ | ○ | ○ |
De novo ligands sized to the pocket Filtered to the cavity heavy-atom window, then docked | ● | ○ | ○ | ◐ |
Approved-drug repositioning panel 3,311 approved drugs plus 381 curated rescue ligands | ● | ○ | ○ | ○ |
Chemical space and developability alerts ECFP4 Tanimoto, scaffold hop, PAINS and Brenk alerts | ● | ○ | ○ | ○ |
| Mechanism and scoring honesty | ||||
Motif attribution for hotspot variants Zinc coordination, L2/L3 and DBD core, from static SAE profiles | ● | ○ | ○ | ◐ |
Rescue-vs-inhibit reading Transparent features with a calibrated uncertainty band | ● | ○ | ○ | ○ |
Size-bias guard on rankings Ligand efficiency, not raw docking score | ● | ○ | ○ | ○ |
Measured read-across affinity with bands 4,500 published affinities, MAE 1.14 pKd at close analogues | ● | ○ | ○ | ○ |
| Evidence and validation | ||||
DMS, ClinVar and cancer cohort context 7,448 DMS variants, 378 ClinVar variants, NCI GDC | ● | ○ | ○ | ◐ |
Prime editing sensor evidence Sánchez-Rivera Lab screen, 1,227 variants | ● | ○ | ○ | ○ |
Live literature check per step Europe PMC verdict for variant, pocket and ligand | ● | ○ | ○ | ○ |
Validated against published rescue data 68 compound–variant pairs across 23 variants | ● | ○ | ○ | ○ |
Suggested next experiment Assay, construct and control for the current shortlist | ● | ○ | ○ | ○ |
| Cell response and export | ||||
Cell pathway response (illustrative) p53/MDM2/p21/Bax model, not a clinical prediction | ● | ○ | ○ | ○ |
Batch upload of variant panels CSV, TSV or Excel through the identical pipeline | ● | ○ | ◐ | ◐ |
Full provenance and reproducible export Inputs, scorer version and sources recorded; null when uncomputed | ● | ◐ | ◐ | ◐ |
● included · ◐ partial or needs extra tools · ○ not in scope. Reflects each tool's core scope as typically used; Oncyra uses ESMFold and OpenFold3 internally.
Full technical capability list (34)Show
Continuous workflow
Variant entry, receptor choice, modelling, docking, pose diligence and shortlisting all live on one screen, so context is never lost between steps.
Interactive 3D residue view
Click a residue in the sequence and the viewer focuses on it with legible atom, residue and element labels at any zoom.
In-browser chemical space
ECFP4 fingerprints computed in a worker and cached locally: Tanimoto search, scaffold-hop view, descriptors with PAINS/Brenk alerts, PCA map and CSV export. Descriptor arithmetic only.
Side-by-side ligand comparison
Compare the docked shortlist in one dialog: 2D structures with MW, cLogP, HBD/HBA, TPSA, rotatable bonds and Ro5 violations.
Ligand ↔ residue interactions
Every dock is followed by a deterministic geometric scan for hydrogen bonds and hydrophobic contacts, residue by residue.
Pose sanity checks
Eight internal pose checks (ligand present, internal clash, protein-ligand clash, inside the receptor, bond lengths, compactness, element sanity, finite coordinates) run on every pose alongside its score.
Modular generative pipeline
Generation and evaluation stay separate. Pocket volume sets a heavy-atom budget for the generator; returned SMILES are re-parsed, re-filtered, embedded and docked by the same local physics.
Physics-first rescue shortlist
The table ranks the library by cavity fit before anything is docked, so novel hits can surface on geometry alone rather than only known names.
Approved-drug repositioning
3,311 approved drugs can be swept against any TP53 pocket in-session: ranked by cavity fit, then the best-fitting batch is docked. A sweep this size under a simplified scorer produces predictable false positives - highly charged or flat molecules that fit in vacuum but fail on permeability or plasma-protein binding - so treat it as a triage shortlist, not a repositioning claim.
Live measured binding
The measured-binding card queries public bioactivity archives live for published Kd, Ki, IC50 and EC50 with assay and citation. Nothing is imputed.
Disease and homology context
Live TP53 disease associations and structurally similar PDB entries sit beside the loaded receptor, with a secondary source for bioactivity.
Medicinal chemistry suite
Paste any hit SMILES to get a synthetic-accessibility score, an extended rule-based ADMET profile (solubility, GI absorption, hERG risk, CYP, P-gp), bioisosteric analogues and a closed lead-optimisation loop ranked on SA, ADMET quality and scaffold similarity.
Retrosynthesis hints
The docked ligand is scanned for named-reaction retrons and every plausible disconnection is listed. Single-step only, not a route search.
NCI GDC cohort context
Live per-project case counts, cohort frequencies and top co-mutated genes for the selected variant. Overlay only — rankings are unchanged.
Mutation evidence explorer
Pocket class, ΔΔG, DMS and sensor readouts, cohort frequency and curated literature gathered into one collapsible card next to the prediction.
Residue overlay track
A 1D sequence track aligns per-residue DMS and prime-editing sensor values, aggregated per codon so hotspot signal is not diluted.
Rescue-vs-inhibit scatter
pRescue against pInhibit for the docked shortlist, separating rescuers, inhibitors and ambiguous compounds before any ranking call.
Live cell physiology (illustrative model)
An in-browser ODE-driven p53/MDM2/p21/Bax network uses Human Protein Atlas cell-line consensus nTPM for the selected context plus literature-derived protein and turnover anchors. It is one generic rescue mechanism applied qualitatively across the catalogue, not variant-specific pathway rewiring: apply a docked hit, vary dose, watch the pathway respond, and export the run as a video — not a clinical prediction.
Europe PMC evidence backbone
Variant, receptor, pocket and every ligand are checked live against the full-text literature index, returning a per-step verdict with the papers behind it.
Batch mode and run history
Queue several variants and re-open any past session: docks, generated structures and designs persist per variant in your browser.
Re-measurable benchmark ledger
Enrichment, calibration, site labelling, cavity detection and pose accuracy can each be re-measured in your own browser, recorded next to the published value.
Manual design lane
Draw a molecule in the built-in editor, save it per variant and dock it through the identical pipeline. Manual designs never enter benchmark counters.
A–E grade on every hit
One letter folds pose quality, site relevance, geometry, efficiency and published evidence into a readable verdict; click it for the full weighted breakdown.
Literature-anchored ranking
Published evidence for a compound against the exact variant or its mechanism class is scored and blended with the pose result, itemised and traceable.
De-novo protein binders
Generate a mini-protein backbone for the mutant pocket, design a sequence at your chosen length and view it beside the receptor, with a foldability pre-check.
Honest small-screen statistics
Runs of three to seven compounds report enrichment with a resampled 95% interval; an interval crossing 1x plainly says the ordering is not yet distinguishable from chance.
On-demand stability prediction
An independent published ΔΔG predictor can be called for any single substitution. Opt-in, measured values win, and an unreachable service is reported as such.
Cavity detection and receptor QC
Pockets are found by an in-browser P2Rank port with a geometric fallback, cached locally; a completeness gate flags missing atoms or chain breaks.
Consensus and ensemble docking
Multiple receptor conformations and scoring passes are compared, agreement is reported and rank-fusion combines them, so setup-specific wins are visible. Predicted receptors come by default from ESMFold, a single-sequence predictor that uses no multiple-sequence alignment, and the consensus card states that plainly so a predicted fold is never mistaken for an evolutionarily-informed one. An alignment-informed route is available as an opt-in: an NVIDIA MSA search feeding OpenFold3, labelled separately wherever it is used.
Suggest the next experiment
For the current shortlist the lab proposes the assay, construct and control that would best discriminate the hypothesis, with the expected outcome.
Batch upload from spreadsheets
Drop a CSV, TSV or Excel panel of variants and each one is queued through the identical pipeline, with results written to run history.
AI/ML data behind the app
Models use public, citable data only: ~374k pose-RMSD measurements, complex-with-affinity records across 8 benchmarks, curated ΔΔG sets, ChEMBL bioactivities and Human Protein Atlas cell-line consensus RNA. Deep-mutational-scanning panels inform descriptive SAE profiles only; validation and scoring rely on an independent stability model, and no held-out benchmark compound is used for fitting.
Reproducible screen export
One click exports the screen as organised JSON, or a full bundle with report, CSV tables, figures and structure files. Content-hashed; uncomputed fields stay null.
Open provenance and audit trail
Every screen records its inputs, scorer version and the exact public sources behind each number, so any result can be retraced and defended end to end.
Powered by measured analogues, not a black box
Docking scores are never converted into an affinity. The only affinity value shown beside a candidate is read across from the nearest measured analogues in a 4,500-compound set of published affinities (HiQBind, BindingMOAD, GatorAffinity p53 records), as a similarity-weighted mean over the three closest molecules above an ECFP4 Tanimoto cutoff, and it is labelled as read-across wherever it appears. On held-out validation the close-analogue tier reaches mean absolute error 1.14 pKd with 80% within ±1.80 pKd, and every value carries an honest conformal ±band. A ligand-only neural predictor was benchmarked on the same held-out set and failed it (Pearson r = -0.23), so it is not used for the reported column.
4,500 affinities
Read-across draws nearest neighbours from published measured affinities (HiQBind, BindingMOAD, GatorAffinity p53 records), not from a model's training set.
MAE ≈ 1.14 pKd
Held-out validation on 800 measured complexes. 80% within ±1.80 pKd at the close-analogue tier (Tanimoto ≥ 0.5, 52% coverage). For the remaining ~48% there is no close measured analogue: those rows show no affinity value and fall back to physics-only ordering. Honest conformal ±bands, not an extrapolation.
No upload, no queue
The ECFP4 similarity search and read-across run client-side — every screened or generated molecule is scored straight from its SMILES string, with nothing sent to a server.
Real numbers, cited sources, one continuous workflow.
These benchmarks measure different capabilities and should not be interpreted as a single overall accuracy score.
Total experimental and computational measurements across 8 public benchmarks (1,361,567 unique complex-with-affinity records + 374,518 pose-RMSD measurements; per-source counts on /rescuebench; each run uses only the subset with a resolvable structure/ligand pair)
STRUCTURAL: Structural Pose Quality. 374,518 experimental complexes; Median heavy-atom RMSD 0.77 Å · 86.7% below 2 Å.
FUNCTIONAL: Functional Variant Effect. 7,448 TP53 DMS variants; Spearman ρ 0.38 (95% CI 0.36–0.40).
CLINICAL: Clinical Variant Classification. 378 ClinVar TP53 variants; 65.1% agreement (95% CI 60.3–69.8%) · sensitivity 0.46, specificity 0.95.
RESCUE: Compound Rescue Evidence. 68 experimental compound-rescue pairs; 76.5% agreement (95% CI 66.2–86.8%) · sensitivity 0.69, specificity 0.85.
Approved drugs (ChEMBL max_phase 4) available for in-browser repositioning docks against any variant
TP53 DBD missense variants in the catalogue, searched live for measured binding data in ChEMBL; the operative subset is the variants with a resolvable query and returned record; most have no published affinity, reported honestly
Curated TP53 rescue ligands from ChEMBL, PubChem, and published catalogues
Full-text life-science articles indexed in Europe PMC, queried live to anchor every pipeline recommendation in published evidence rather than opaque scoring
TP53 DBD missense variants passing the structural dockability gate, out of 4,137 missense variants in the catalogue
Strict binary benchmark: 10 variants · F1 0.857 · in-sample fit check. The rescue evidence above and this strict benchmark are scored separately and never merged. Every pair is listed on the benchmarks page.
What the workspace helps a p53 biologist do
A single screen carries a p53 variant from selection to a scored, evidence-linked shortlist. The per-mutation view sits next to experimental data and citations, and every visible result is exportable as a single reproducible snapshot.
Deterministic physics core
Structure viewing, ligand geometry, docking, and scoring run through a transparent UFF Lennard-Jones plus hydrogen-bond force field with a distance-dependent dielectric. No opaque model file, no hidden calibration.
Prediction, on demand
When a mutation has no suitable experimental structure, ESMFold generates a mutation-aware model and streams it back to the viewer across the 4,137 TP53 DBD missense variants in the catalogue. ESMFold is single-sequence: it uses no multiple-sequence alignment, so every predicted fold is labelled as such, with its confidence reported on the standard 0-100 scale. An alignment-informed OpenFold3 route, fed by an NVIDIA MSA search, is available as an opt-in and is labelled separately.
Provenance you can filter
Every card carries a provenance tag - measured, measured-modelled or derived - and a global toggle can withhold everything that is not a wet-lab measurement. Dataset row counts, molecule coverage and validation checks are published on the data-quality page.
From a variant to an inspectable 3D scene, with every pose checked before it is drawn.
Mutation-aware structures
4,137 variantsESMFold models when no experimental PDB fits the variant, across the 4,137 TP53 DBD missense variants in the catalogue. Each is labelled single-sequence, with confidence on the 0-100 scale.
Interactive residue focus
Click any sequence position to zoom the 3D viewer, with legible atom, residue and element labels at any scale.
Chemistry-accurate rendering
CPK element colours and van der Waals radii, single, double, triple and aromatic bond geometry, wedge and dash stereo cues, and a ligand-versus-protein scale control. One compound in the scene by default.
Pose gate and interaction map
8 pose checksA structure-validation gate runs before render, with the eight internal pose checks and a per-pose ligand-residue interaction map.
Crystal-ligand overlay
Docking visualisation panel showing pose inputs, residue-numbering offsets and ligand provenance, with centroid offset and per-atom distance measured against every crystal ligand in the receptor file.
Look up a variant. Get catalogue-backed records.
Variant records come from deterministic lookup over the curated mutation catalogue, so every figure maps back to a catalogue row with its citation. A hosted language model sits on top for semantic search and literature synthesis, and its prose is kept separate from the catalogue-backed records.
>Curated rescue compounds for R248Q
>TP53 hotspots ranked by dockable-pocket score
>R175H versus R273H mechanism records
Frequently asked, honestly answered.
What it does, how confident it is, and where it stops. Ten chapters, plain answers. Pick one to begin.
Open a variant and work through it end-to-end.
Sign in with your credentials. Accounts are provisioned by the Oncyra team.